Entropy fingerprint
EntropyFingerprint gives the pair entropy fingerprint of each atom, a
measure of local order that needs no reference structure: low (more
negative) in crystals, high in liquids and gases.
import numpy as np
import mdapy as mp
from mdapy.analysis import EntropyFingerprint
from mdapy.build import build_crystal

# fcc copper of 4 x 4 x 4 cubic cells, slightly disordered
crystal = build_crystal("fcc", 3.615, elements="Cu", repeat=(4, 4, 4))
shake = np.random.default_rng(0).normal(0, 0.05, (len(crystal), 3))
system = crystal.with_columns(positions=crystal["positions"] + shake)

result = EntropyFingerprint(cutoff=5.0, sigma=0.2, average_cutoff=3.0).compute(system)
result.entropy  # in units of the Boltzmann constant
system = system.with_columns(result)  # the column entropy_average
In a polycrystal, cut in half:
from mdapy.build import build_crystal, build_polycrystal
from mdapy.visualization import render

grains = build_polycrystal(
    build_crystal("fcc", 3.615, elements="Cu"), mp.Box(50.0), 5, seed=4
)
fingerprint = EntropyFingerprint(cutoff=5.0, sigma=0.2, average_cutoff=3.0).compute(
    grains
)
render(grains, "images/entropy.png", atoms=grains["positions"][:, 0] < 25,
       color=fingerprint.entropy, width=800, height=600, legend="entropy")  # fmt: skip
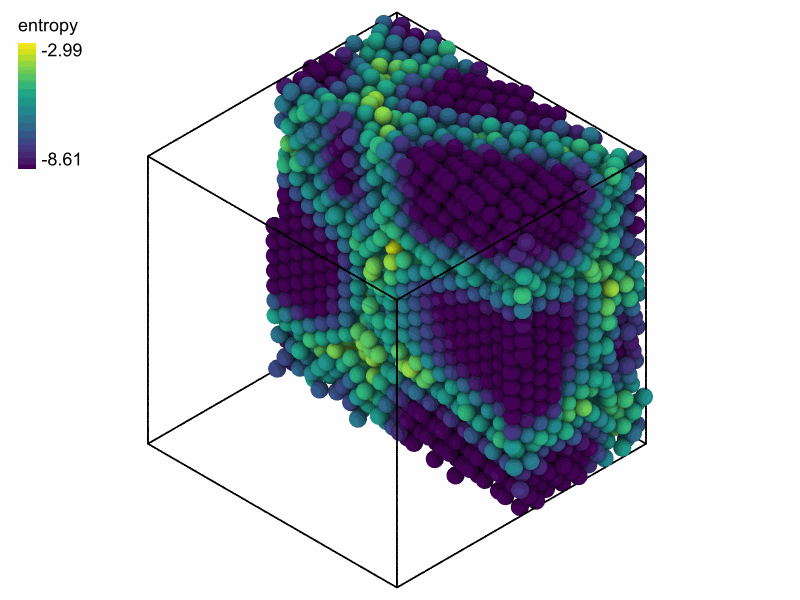
The averaged entropy is lowest in the ordered grains (dark) and highest at the boundaries and their junctions (green and yellow).
Parameters
| Name | Type | Default | Unit | Meaning |
|---|---|---|---|---|
cutoff |
float64 | Å | neighbors are closer than it | |
sigma |
float64 | 0.2 | Å | width of the Gaussian of each neighbor, below cutoff |
local_density |
bool | False |
use the density of each atom's neighbors instead of that of the system | |
average_cutoff |
float64 | None |
Å | average each value with those of the neighbors closer than it |
compute(system, *, neighbors=None, progress=None, cancel=None) takes the
arguments of every analysis. A given neighbor list
must reach the larger of cutoff and average_cutoff; without one, the
neighbors within cutoff are found and used at once, never stored.
Results
An EntropyResult:
| Attribute | Type | Meaning |
|---|---|---|
entropy |
(N,) float64 | the fingerprint of each atom, in units of the Boltzmann constant; its column entropy, with the options in the name: entropy_average, entropy_local_density, entropy_average_local_density |
average |
bool | whether the values are averaged |
Errors
The constructor raises InvalidParameter for a cutoff that is not
positive, a sigma not between 0 and the cutoff, or an average_cutoff
that is not positive; compute raises it for a neighbor list that does not
fit.
Definition
For each atom \(i\) a local radial distribution is built from its neighbors \(j\) closer than the cutoff \(r_c\), each smeared by a Gaussian of width \(\sigma\):
where \(\rho\) is the number density of the system or, with local_density,
the number of neighbors over the volume of the cutoff sphere. \(g_i\) is
evaluated at \(r = k\sigma\) up to the cutoff and the integral is the
trapezoid rule over those points, as in LAMMPS; where \(g_i < 10^{-10}\) the
integrand is \(r^2\). With average_cutoff, \(s_i\) is averaged with the values
of the neighbors closer than it.
LAMMPS compute entropy/atom spreads each neighbor over seven points only;
MDAPY uses the whole Gaussian, so that equivalent atoms get equal values.
The two agree to about 0.5 %.
See also
- LAMMPS
compute entropy/atom.
Please cite