Molecular dynamics
mdapy.simulation.MolecularDynamics(system, potential, *, timestep=0.001)
runs molecular dynamics of a system in LAMMPS, with the interaction of a
LAMMPS calculator, and gives back the thermodynamic state, the
atoms along the way and the system at the end.
from mdapy.build import build_crystal
from mdapy.calculators import LAMMPS
from mdapy.simulation import MolecularDynamics

potential = LAMMPS(
    "pair_style eam/alloy\npair_coeff * * Cu_mishin1.eam.alloy Cu", elements=["Cu"]
)
crystal = build_crystal("fcc", 3.615, elements="Cu", repeat=(8, 8, 8))

with MolecularDynamics(crystal, potential, timestep=0.002) as md:
    md.velocities(600, seed=1)
    md.run(10000, ensemble="npt", temperature=300, pressure=0.0, thermo=100)
    md.run(5000, ensemble="nvt", temperature=300, frames=500)
    md.thermo  # polars DataFrame: step, time, temperature, energies, ...
    md.frames  # the atoms every 500 steps of the second run
    hot = md.system  # the atoms at the end, with velocities and images
The thermodynamic state along the two runs:
from mdapy.visualization import plot

thermo = md.thermo
a = (thermo["volume"] / 512) ** (1 / 3)  # lattice parameter of the 8 x 8 x 8 cells
with plot.style():
    fig, (top, bottom) = plot.figure("single", 100, nrows=2, sharex=True)
    top.plot(thermo["time"], thermo["temperature"])
    top.set_ylabel("Temperature (K)")
    bottom.plot(thermo["time"], a)
    bottom.set_ylabel("Lattice parameter (Å)")
    bottom.set_xlabel("Time (ps)")
    for ax in (top, bottom):
        ax.axvline(20.0, color="0.5", linestyle="--")  # npt, then nvt
    plot.save(fig, "images/dynamics", formats=("png",))
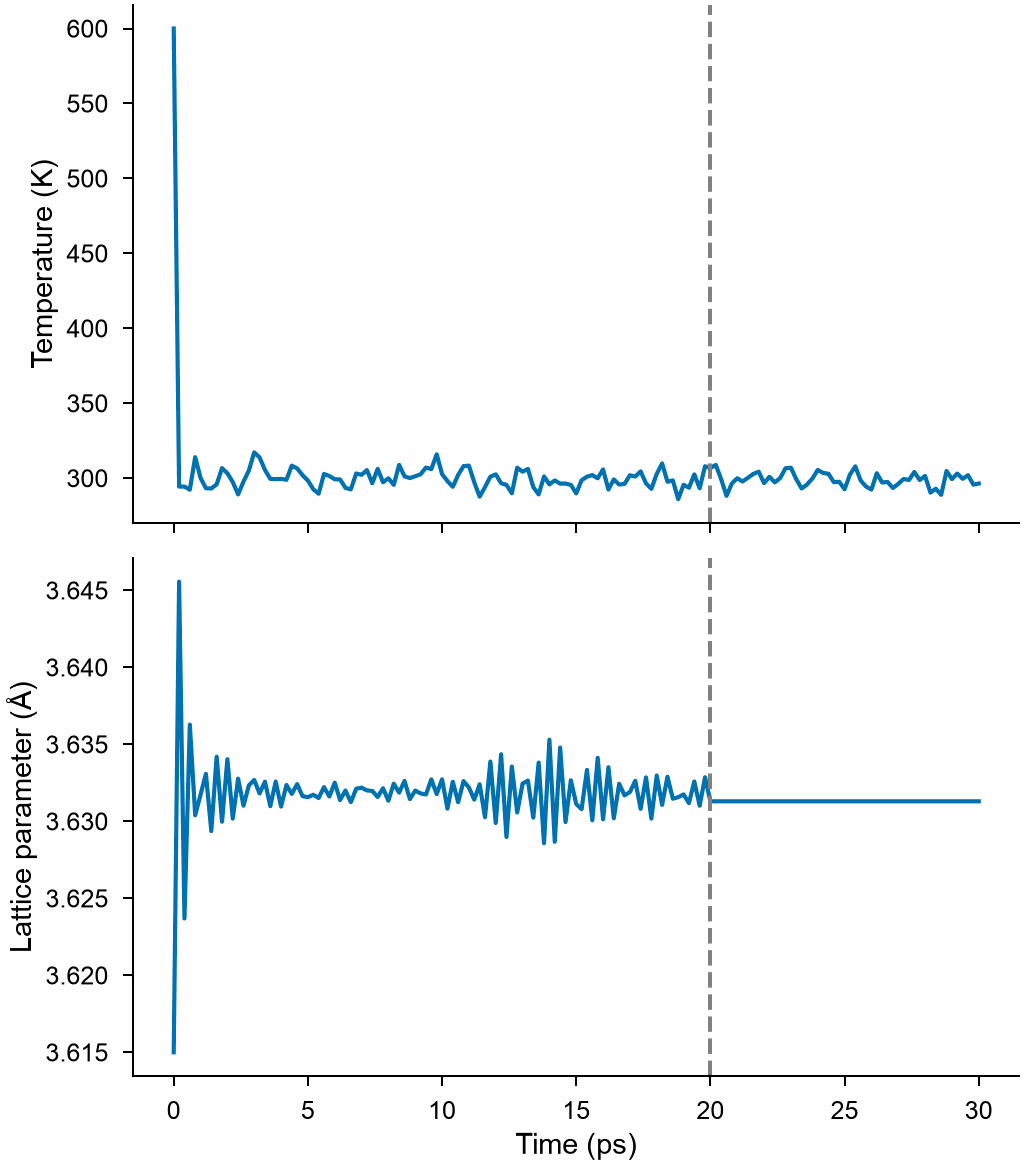
Copper started at 600 K: half the kinetic energy goes into the potential energy at once, and the thermostat holds the rest at 300 K. The barostat expands the box from the lattice parameter of 0 K to that of 300 K at zero pressure; after the dashed line the box is fixed (nvt).
LAMMPS runs in metal units, those of MDAPY, through its Python module,
installed as the LAMMPS documentation
describes; potential gives the commands of the interaction, the element
of each atom type and the command-line arguments of LAMMPS, such as those
of Kokkos. The box may have any orientation and need not be periodic:
LAMMPS gets it rotated into its own form, its non-periodic directions
shrink-wrapped but never smaller than the box of system, and everything comes
back in the orientation of system.
The velocities of system, if it has them, are where the dynamics starts.
| Name | Type | Default | Unit | Meaning |
|---|---|---|---|---|
system |
System |
the atoms, with an element column, and velocities in Å/ps when it has them |
||
potential |
LAMMPS |
the interaction, as a LAMMPS calculator | ||
timestep |
float64 | 0.001 | ps | the time step |
Running
| Method or attribute | Meaning |
|---|---|
velocities(temperature, *, seed) |
random velocities of a temperature in K, Gaussian, with no total momentum; the same for the same seed and atoms, whatever their positions |
run(steps, *, ensemble="nve", temperature=None, pressure=None, barostat="iso", thermostat_damping=None, barostat_damping=None, thermo=100, frames=None, progress=None, cancel=None) |
runs steps steps |
command(commands) |
runs any LAMMPS commands, one or several lines |
lammps |
the LAMMPS instance itself |
close() |
closes the instance; leaving a with block closes it too |
Parameter of run |
Type | Default | Unit | Meaning |
|---|---|---|---|---|
ensemble |
str | "nve" |
"nve", or "nvt" with a Nosé–Hoover thermostat, or "npt" with a thermostat and a barostat, or "nph" with a barostat alone (LAMMPS fix nve, nvt, npt, nph) |
|
temperature |
float64 or (float64, float64) | K | the temperature of the thermostat, or its values at the start and end of the run; for "nvt" and "npt" |
|
pressure |
float64 or (float64, float64) | GPa | the pressure of the barostat, or its values at the start and end; for "npt" and "nph" |
|
barostat |
str | "iso" |
the box changes alike in every direction ("iso"), along each axis ("aniso"), in any way ("tri"), or alike along its first two vectors with the third kept ("xy", as for a sheet) |
|
thermostat_damping |
float64 | 100 time steps | ps | the damping time of the thermostat |
barostat_damping |
float64 | 1000 time steps | ps | the damping time of the barostat |
thermo |
int32 | 100 | records the thermodynamic state every this many steps, and at the start and the end | |
frames |
int32 | None |
keeps the atoms every this many steps | |
progress |
callable | None |
called as progress(done, total) with the steps run |
|
cancel |
CancelToken |
None |
stops the run, which raises Cancelled |
A run is a LAMMPS run in pieces between the steps at which something is
recorded; the pieces run as one run would, a temperature or pressure
changing from its start to its end value across all of them.
Results
| Attribute | Type | Meaning |
|---|---|---|
system |
System |
the atoms now, with velocities and image columns, the box, and timestep and time in info; the other columns of system, but forces and energies, which are out of date |
thermo |
polars DataFrame |
the thermodynamic state of all the runs so far, one row per record |
frames |
tuple of System |
the atoms kept in all the runs so far, as system gives them |
step |
int64 | the time step number of LAMMPS |
Column of thermo |
Type | Unit | Meaning |
|---|---|---|---|
step |
int64 | time step number | |
time |
float64 | ps | time |
temperature |
float64 | K | temperature |
potential_energy, kinetic_energy, total_energy |
float64 | eV | energies of all the atoms |
pressure |
float64 | GPa | pressure |
volume |
float64 | ų | volume of the box |
stress |
(6,) float64 | GPa | stress in Voigt order xx, yy, zz, yz, xz, xy, positive under tension, in the axes of system |
Errors
MolecularDynamics raises InvalidParameter for a potential that is not a
LAMMPS calculator, a time step that is not positive, elements without an
atom type, and commands that LAMMPS rejects; MissingProperty for a system
without elements; ImportError without the LAMMPS Python module.
velocities raises InvalidParameter for a negative temperature or a seed
that is not positive; run for a negative number of steps, an unknown
ensemble or barostat, a thermostat without a temperature, a barostat
without a pressure, "aniso" or "tri" in a box that is not periodic,
"xy" in a box not periodic along its first two vectors, and thermo or frames that are not positive; command for
commands LAMMPS rejects; and lammps after close.